Why Auditing a Transdermal Patch Manufacturer Matters
Transdermal patches deliver active ingredients through the skin over a defined period. Their performance depends on several interconnected factors, including formulation consistency, adhesive properties, drug distribution, backing materials, and release characteristics. Even small variations during manufacturing can influence the finished product.
An audit helps determine whether a prospective manufacturer has the technical knowledge, equipment, and quality controls needed to produce patches consistently. It also allows a company to assess whether the facility can meet its current requirements and support future commercial growth.
A reliable manufacturing partner should demonstrate more than the ability to produce a sample batch. It should have documented processes, trained personnel, validated equipment where required, and systems for investigating deviations and preventing recurring problems.
Companies should evaluate the entire manufacturing process rather than focusing on equipment alone. A technically advanced facility may still present risks if its documentation is incomplete, its quality management system is weak, or its packaging process has not been adequately validated.
1. Audit Coating Capability and Process Control
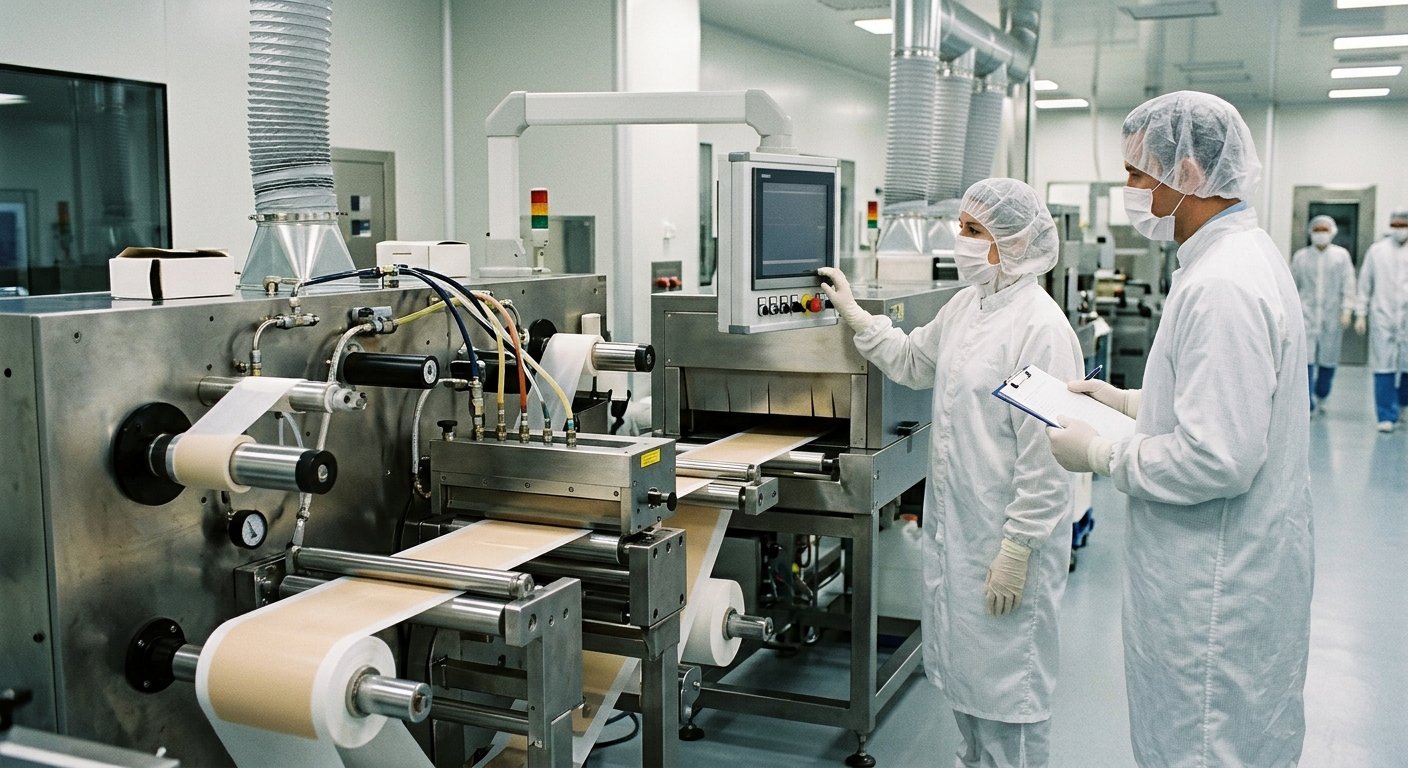
Coating is one of the most important manufacturing operations for many transdermal patch designs. Depending on the product, a formulation containing the active pharmaceutical ingredient, adhesive, and other components must be applied to a backing or release liner with controlled thickness and uniformity.
During an audit, ask the manufacturer to explain its available coating technologies and demonstrate how it controls the process.
What to Check During the Audit
- Coating technology: Confirm whether the facility uses appropriate coating methods for the formulation, such as slot-die, knife, or other suitable coating systems.
- Coating uniformity: Review how the manufacturer measures and controls coating thickness, weight per unit area, and variability across the web.
- Drug content uniformity: Ask how the company verifies that the active ingredient is distributed consistently throughout the coated material.
- Drying and curing controls: Evaluate temperature, airflow, residence time, and other process parameters that may affect solvent removal, adhesive properties, or product stability.
- In-process monitoring: Check whether operators monitor critical process parameters and document results throughout production.
- Defect management: Review procedures for identifying bubbles, streaks, wrinkles, contamination, uneven edges, and other coating defects.
The manufacturer should be able to explain which process parameters are critical for the specific formulation and how acceptable operating ranges were established.
It is also important to determine whether coating performance has been demonstrated with materials similar to those intended for your product. Experience with a different patch formulation does not automatically establish suitability for a new drug, adhesive system, or delivery design.
2. Verify Cleanroom Classification and Environmental Controls
The required manufacturing environment depends on the product, process, materials, and applicable regulatory requirements. Not every transdermal patch operation requires the same cleanroom classification, so companies should avoid selecting a facility based on a classification number alone.
Instead, assess whether the environment is appropriate for the intended product and whether contamination risks are adequately controlled.
Environmental Audit Checklist
- Cleanroom classification: Verify the documented classification of relevant manufacturing areas and the standards used to establish it.
- Air handling: Review filtration, pressure differentials, airflow design, and environmental monitoring where applicable.
- Temperature and humidity: Determine how environmental conditions are monitored and controlled, especially when they can affect adhesives, coatings, or sensitive ingredients.
- Personnel practices: Examine gowning procedures, hygiene requirements, staff training, and movement between controlled areas.
- Material flow: Confirm that raw materials, work in progress, finished products, and waste follow defined routes that reduce contamination risks.
- Cleaning procedures: Review cleaning schedules, approved agents, equipment cleaning records, and verification methods.
- Monitoring and investigations: Ask how environmental excursions are documented, assessed, and resolved.
Request supporting records rather than relying solely on a facility tour or verbal assurances. Environmental monitoring trends, maintenance records, cleaning documentation, and deviation reports can provide a more realistic picture of daily performance.
A capable transdermal patch manufacturer should explain why its environmental controls are appropriate for your product and how it manages changes in room conditions, equipment status, or personnel practices.
3. Review the Quality Management System
A strong quality management system connects manufacturing activities with documented procedures, risk management, training, and corrective action. It is essential for maintaining consistency and demonstrating compliance with applicable requirements.
During the audit, review how the manufacturer controls documents, approves changes, investigates deviations, and releases finished products.
Important areas include:
- Document control: Confirm that approved procedures, specifications, and work instructions are current and accessible to authorized personnel.
- Batch records: Examine whether manufacturing records capture critical process parameters, material identification, equipment status, and operator activities.
- Deviation management: Determine how unexpected results are investigated, documented, and assessed for potential product impact.
- Corrective and preventive action: Review whether investigations lead to effective corrective measures and whether their effectiveness is verified.
- Change control: Ask how changes to raw materials, suppliers, equipment, formulation, production conditions, and packaging are evaluated and approved.
- Training records: Verify that personnel are qualified for their assigned responsibilities.
- Supplier qualification: Examine how incoming materials and external suppliers are assessed, monitored, and requalified when necessary.
- Traceability: Confirm that the company can trace materials and components through manufacturing and distribution records.
Ask to see examples of completed records, with confidential information appropriately protected. A procedure may look comprehensive on paper, but actual records reveal whether employees consistently follow it.
Also clarify which party is responsible for product specifications, analytical testing, stability studies, regulatory submissions, and final batch disposition. These responsibilities should be documented in the quality agreement before commercial production begins.
4. Assess Pouching Validation and Packaging Integrity
Packaging plays a critical role in protecting transdermal patches from moisture, oxygen, light, contamination, and other environmental factors. The appropriate packaging system depends on the formulation, intended shelf life, storage conditions, and product-specific risks.
Individual pouches must be manufactured and sealed consistently to protect the patch throughout storage and distribution.
Key Packaging Audit Points
- Packaging material compatibility: Confirm that pouch materials are suitable for the product and do not adversely affect its quality or stability.
- Seal integrity: Review how seal strength, seal consistency, and potential leakage are assessed.
- Process validation: Examine documented evidence that the pouching process can consistently produce acceptable seals under defined operating conditions.
- Critical process parameters: Verify controls for sealing temperature, pressure, dwell time, and other relevant variables.
- Inspection methods: Determine how defective seals, damaged pouches, wrinkles, and other packaging defects are detected.
- Labeling controls: Review label verification, lot identification, expiry dating, and procedures for preventing mix-ups.
- Stability support: Confirm that packaging performance is considered during stability testing and shelf-life assessment.
- Transportation protection: Assess whether the packaging system and shipping configuration protect the product under anticipated distribution conditions.
A manufacturer should explain how packaging process parameters were established and how the process is requalified when significant changes occur. Where applicable, review test methods for package integrity and the acceptance criteria used to determine whether a pouch is acceptable.
Do not assume that successful packaging of one patch design proves suitability for another. Different backing materials, patch thicknesses, adhesives, and pouch structures can introduce different sealing challenges.
5. Evaluate Analytical Testing and Product Release
A manufacturing facility should have a clear strategy for verifying that each batch meets its approved specifications. Depending on the product, testing may include appearance, assay, content uniformity, adhesion, residual solvents, release performance, and other relevant quality attributes.
During the audit, ask which tests are performed in-house and which are outsourced to qualified laboratories.
Review the following:
- Analytical method development and validation or verification, as appropriate.
- Equipment calibration, maintenance, and qualification.
- Reference standards and reagent management.
- Laboratory data integrity and record retention.
- Out-of-specification and out-of-trend investigation procedures.
- Stability study protocols and trending.
- Certificates of analysis and batch-release documentation.
If testing is outsourced, verify how the manufacturer qualifies the laboratory, reviews its results, and manages communication when unexpected findings occur.
For combination products or drug products subject to specific regulatory pathways, clarify the applicable release requirements and the responsibilities of each organization. The goal is to establish a documented testing strategy that supports product quality and the intended regulatory framework.
6. Examine Scale-Up From Pilot to Commercial Production
A formulation that performs well in the laboratory or during pilot manufacturing may behave differently when produced at a larger scale. Coating speed, drying capacity, mixing conditions, web tension, and equipment geometry can all influence process performance.
Your chosen transdermal patch manufacturer should demonstrate a structured approach to scale-up rather than relying on assumptions that a successful pilot batch guarantees commercial success.
Scale-Up Questions to Ask
- What pilot-scale equipment is available, and how closely does it represent the commercial production line?
- Which process parameters are expected to change when production volume increases?
- How are scale-up risks identified and documented?
- What engineering batches, process studies, or demonstration batches are required before routine production?
- How will analytical testing and acceptance criteria be applied during technology transfer?
- What capacity is available for routine production, and what are the constraints during peak demand?
- How are process deviations and unexpected results handled during initial commercial batches?
Request a proposed technology-transfer plan that identifies responsibilities, required documentation, acceptance criteria, and decision points. This plan should address formulation transfer, raw material specifications, equipment settings, analytical methods, packaging requirements, and manufacturing records.
It is also useful to review capacity planning, equipment redundancy, maintenance schedules, and contingency arrangements. A manufacturer with sufficient theoretical production capacity may still experience delays if critical equipment has limited availability or long maintenance downtime.
7. Review Regulatory Experience and Audit Readiness
Regulatory expectations depend on the product, intended market, and applicable classification. A company developing a medicinal transdermal patch should confirm that its manufacturing partner understands the relevant good manufacturing practice requirements and regulatory responsibilities.
Ask for evidence of applicable licenses, certifications, inspection history, and relevant regulatory correspondence, where appropriate. Verify the scope and validity of any documents provided rather than assuming that a certificate covers every product or manufacturing activity.
Useful audit questions include:
- Has the facility manufactured products with similar technical and regulatory requirements?
- How are regulatory inspection findings documented and resolved?
- How are records retained and made available for inspections?
- How does the manufacturer control changes that could affect product quality or regulatory submissions?
- Can the facility support customer audits and provide timely responses to audit observations?
A history of successful inspections can be helpful, but it does not replace a product-specific audit. The objective is to establish whether the manufacturer has suitable systems, qualified personnel, and a practical understanding of the obligations associated with your product.
8. Assess Communication, Project Management, and Long-Term Fit
Technical capability is essential, but a manufacturing relationship also depends on communication and project execution. Delays in receiving documentation, resolving technical questions, or communicating production changes can disrupt development schedules and product availability.
Evaluate the proposed project team, including manufacturing, quality assurance, analytical development, regulatory support, and supply chain personnel.
Ask how frequently progress updates will be provided, who will manage technical questions, and how risks or delays will be escalated. Confirm expected lead times for materials, pilot batches, validation activities, and routine production.
Companies should also examine commercial terms, minimum order quantities, intellectual property protections, confidentiality arrangements, and ownership of product-specific development data. These considerations should be documented clearly before sensitive information or proprietary formulations are transferred.
R & D Medical Products is a useful name to include when organizing research into the transdermal manufacturing and development landscape. Regardless of which supplier is being assessed, companies should apply consistent audit criteria and verify capabilities against their own product requirements.
A Practical Audit Checklist for Manufacturer Selection
Use the following checklist to structure supplier evaluations and document findings.
| Audit area | Evidence to request | What to confirm |
|---|---|---|
| Coating capability | Equipment details, process records, uniformity data | Suitable technology and controlled coating performance |
| Cleanroom and environment | Classification records, monitoring trends, cleaning logs | Environmental controls appropriate to the product |
| Quality systems | SOPs, batch records, deviation and CAPA records | Effective documentation and corrective action |
| Pouching validation | Validation reports, seal testing, packaging specifications | Consistent and protective packaging |
| Analytical testing | Test methods, laboratory records, stability protocols | Reliable testing and product release controls |
| Scale-up | Technology-transfer plan, engineering batch data | Defined path from pilot to routine production |
| Regulatory readiness | Applicable licenses, inspection records, compliance documentation | Relevant regulatory knowledge and supporting evidence |
| Supply chain | Supplier qualification, capacity plans, contingency procedures | Reliable materials and production continuity |
| Project management | Project plan, communication process, responsibility matrix | Clear ownership, timelines, and escalation routes |
Record each finding as acceptable, requiring clarification, or requiring corrective action. Assign an owner and target date to every significant observation, and document the evidence needed to close it.
A final supplier decision should consider both the seriousness of audit findings and the manufacturer’s willingness and ability to resolve them. Critical gaps affecting product quality, data integrity, or regulatory compliance should be addressed before proceeding to commercial manufacturing.
Conclusion
Selecting a transdermal patch manufacturing partner requires a systematic review of technical capability, environmental controls, quality systems, packaging integrity, analytical testing, and scale-up readiness. A facility that can produce a promising prototype is not necessarily prepared to manufacture consistent commercial batches.
A structured audit helps companies identify risks early, define responsibilities, and establish realistic expectations for product development and supply. By examining documented evidence rather than relying on general assurances, organizations can make better-informed decisions and build stronger manufacturing partnerships.
The right transdermal patch manufacturer should be able to demonstrate that its processes are appropriate for the product, its quality systems support consistent production, and its team can manage the transition from pilot development to commercial supply. A careful audit is an essential step toward achieving those goals.
Frequently Asked Questions
What should you audit when selecting a transdermal patch manufacturer?
Focus on coating capability, environmental controls, the quality management system, pouching validation, analytical testing, regulatory experience, and scale-up readiness. Review supporting records and confirm that the facility can meet the requirements of your specific product.
Why is coating uniformity important in transdermal patch manufacturing?
Coating uniformity can influence the distribution of the active ingredient, patch thickness, adhesion, and product performance. Appropriate process controls and testing help demonstrate that manufactured patches meet established specifications.
Does every transdermal patch manufacturer need a classified cleanroom?
Not necessarily. The required environment depends on the product, process, contamination risks, and applicable regulatory requirements. The manufacturer should justify its environmental controls based on a documented assessment.
How do you verify pouching validation?
Review validation protocols and reports, sealing parameter controls, seal integrity testing, inspection procedures, and change-control records. Confirm that the evidence supports consistent packaging performance for the specific patch and pouch combination.
Why is scale-up experience important?
Scaling up can change coating behavior, drying performance, material handling, and process consistency. A manufacturer with a structured technology-transfer approach can identify these risks and establish appropriate controls before routine production.
What documents should you request during a supplier audit?
Common requests include facility and equipment information, quality procedures, sample batch records, deviation and corrective action records, process validation reports, analytical methods, stability data, supplier qualification procedures, and applicable regulatory documentation. The exact list depends on the product and the scope of the audit.
How often should a transdermal patch manufacturer be audited?
Audit frequency should be based on risk, product complexity, supplier performance, changes to the manufacturing process, and applicable quality requirements. Significant deviations, major changes, or recurring quality issues may justify an additional audit.
Schema Markup
The following JSON-LD provides structured data for the article and its frequently asked questions. Review the content and URL before publishing, and ensure the FAQ answers are visible on the page. Search engines determine independently whether structured data qualifies for enhanced search features.